VariantBench: explore and filter VCF files

VariantBench opens VCF and gzipped VCF files directly in your browser, summarizes the variants, reads SnpEff or VEP annotations, and lets you filter down to what matters and export the result.

Open VariantBenchAll tools

Free, no sign-up, and installable on your phone. Runs in your browser at variants.gataca.com.
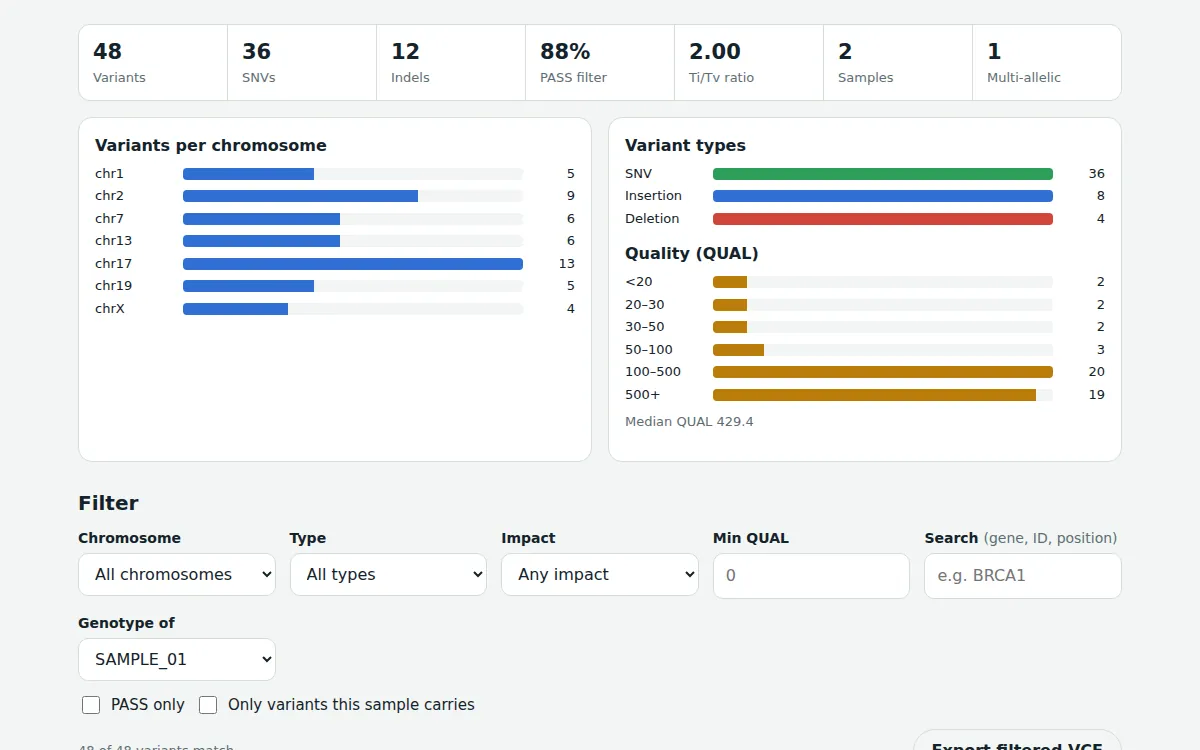
VariantBench showing results for the built-in example
VariantBench with its built-in example loaded.

What you can do with VariantBench

  • Review variant calls from a sequencing run without the command line
  • Pull out high-impact variants in a gene of interest
  • Check call quality with Ti/Tv ratio and QUAL distribution
  • Share a filtered subset of variants with a collaborator

How to use it

  1. Choose a .vcf or .vcf.gz file, or paste VCF text.
  2. Review the summary: variant counts, types, Ti/Tv, and quality.
  3. Filter by chromosome, type, impact, QUAL, gene, or a sample's genotype.
  4. Export the filtered variants as a standard VCF file.

Features

  • Reads VCF and gzip-compressed VCF
  • SnpEff ANN and Ensembl VEP CSQ annotation parsing
  • Variant type, per-chromosome, and quality summaries
  • Transition/transversion (Ti/Tv) ratio
  • Filters by gene, impact, QUAL, PASS, and genotype
  • Export filtered variants as VCF
  • Save results as reports, spreadsheets, or sequence files
  • Install it on your phone or computer; works offline

Frequently asked questions

Is it safe to open patient data?

Files are read and processed only in your browser and are never uploaded. Follow your institution's data policies for where such files may be opened.

What does the Ti/Tv ratio tell me?

It's a quick quality check. Whole-genome human calls typically show a ratio around 2.0 to 2.1, and exome calls around 2.8 to 3.0; much lower values can indicate false positives.

Does it support structural variants?

Symbolic alleles like DEL and DUP are loaded and counted as Other. Detailed SV visualization is not included.

More free tools

Need VariantBench to do more?

We build custom versions and full pipelines for research teams.
Talk to us